Epigenetics
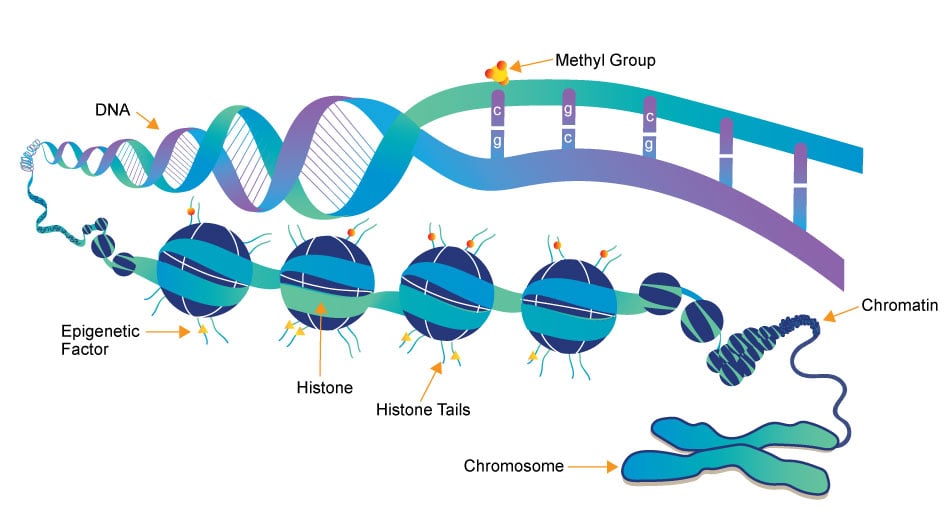
Epigenetics research delves into the molecular mechanisms that control gene expression and cellular traits without altering the underlying DNA sequence. One crucial aspect of this field is the role of small molecules, which act as powerful regulators of epigenetic modifications. These small compounds, typically comprising a few dozen to a few hundred atoms, have emerged as essential tools in understanding and manipulating the epigenome.
- DNA Methylation Inhibitors: Small molecules like 5-azacytidine and 5-aza-2'-deoxycytidine are DNA methyltransferase inhibitors. They block the addition of methyl groups to DNA, leading to DNA demethylation. This can reactivate silenced genes, potentially offering therapeutic avenues for conditions like cancer.
- HDAC inhibitors: HDACs remove acetyl groups from histone proteins, contributing to gene repression. Small molecule HDAC inhibitors, such as Vorinostat and Romidepsin, can reverse this process by increasing histone acetylation, allowing genes to be more accessible for transcription. These inhibitors are being explored for cancer therapy and other conditions.
- Histone Methyltransferase Inhibitors: Small molecules like GSK126 inhibit specific histone methyltransferases, affecting histone methylation patterns. This can alter gene expression, making them promising candidates for cancer and other diseases with epigenetic dysregulation.
- RNA Modulators: Small molecules can also target non-coding RNAs involved in epigenetic regulation. For instance, small molecules called small interfering RNAs (siRNAs) can be designed to target and degrade specific long non-coding RNAs, influencing gene expression.
- Epigenetic Reader Domain Inhibitors: These small molecules target proteins that recognize and bind to specific epigenetic marks. Examples include inhibitors of bromodomain-containing proteins (BET inhibitors), which can disrupt gene regulation by interfering with protein-DNA interactions.
Small molecules in epigenetics research not only provide insights into the fundamental biology of gene regulation but also hold immense promise for developing novel therapeutics. Their ability to selectively modulate specific epigenetic marks and pathways has led to ongoing clinical trials and drug development efforts for various diseases, including cancer, neurological disorders, and inflammatory conditions. Understanding and harnessing the power of these small molecules is at the forefront of modern epigenetics research, offering new hope for precision medicine and targeted therapies.
3 key components involved in the regulation of epigenetic modifications
Epigenetics Writer
Epigenetics writers are enzymes responsible for adding chemical marks or modifications to DNA or histone proteins. These marks include DNA methylation (addition of methyl groups to DNA) and histone modifications (such as acetylation, methylation, phosphorylation, etc.).
Epigenetics Reader
Function: Epigenetics readers are proteins that can recognize and bind to specific epigenetic marks on DNA or histones. These reader proteins interpret the epigenetic code and facilitate downstream cellular processes, such as gene activation or repression.
Epigenetics Eraser
Function: Epigenetics erasers are enzymes responsible for removing or reversing epigenetic marks on DNA or histones. This process allows for the dynamic regulation of gene expression and the resetting of epigenetic states during various stages of development and in response to environmental changes.
-
HDAC1/2 and CDK2 Inhibitor
HDAC1/2 and CDK2-IN-1 is a dual inhibitor targeting HDAC1, HDAC2, and CDK2, with IC50 values of 70.7 μM, 23.1 μM, and 0.80 μM, respectively. This compound effectively disrupts the cell cycle and promotes apoptosis in tumor cells, demonstrating significant in vivo antitumor activity. It is suitable for research applications focused on cancer biology and therapeutic interventions targeting histone deacetylases and cyclin-dependent kinases. -
HDAC/MBLAC2 Inhibitor
Pracinostat dihydrochloride is a potent inhibitor of histone deacetylases (HDACs), demonstrating IC50 values in the range of 40-140 nM, making it a valuable tool in cancer research. In addition, it effectively inhibits metallo-β-lactamase domain-containing protein 2 (MBLAC2) with an EC50 below 10 nM, highlighting its potential use in studies related to epigenetic regulation and resistance mechanisms in cancer therapies. -
PAD1 Inhibitor
D-Cl-amidine is a selective inhibitor of the peptidylarginine deiminase 1 (PAD1) enzyme. It exhibits significant biological activity in modulating citrullination processes involved in various physiological and pathological conditions. This reagent is primarily utilized in research applications focusing on inflammatory diseases, neurodegeneration, and cancer biology, providing essential insights into the role of PAD1 in cellular functions. D-Cl-amidine demonstrates a favorable toxicity profile, supporting its use in experimental settings. -
Aurora B Inhibitor
Barasertib dihydrochloride is a selective inhibitor of Aurora B kinase, exhibiting an IC50 of 0.37 nM in cell-free assays. This compound effectively induces growth arrest and apoptosis in various cancer cell lines, making it a valuable tool for cancer research. Its mechanism of action provides insights into the role of Aurora B in cell cycle regulation and tumorigenesis. -
JAK1 Inhibitor
Ivarmacitinib sulfate is a selective inhibitor of the Janus kinase 1 (JAK1) pathway, exhibiting a significant preference over JAK2, JAK3, and Tyk2. This compound effectively inhibits JAK1-STAT3 phosphorylation, leading to the apoptosis of hepatic stellate cells. Ivarmacitinib sulfate demonstrates notable anti-proliferative and anti-inflammatory properties, making it a valuable tool for research on hepatic diseases and inflammation-related disorders. -
Aurora/JAK Inhibitor
AT9283 lactic acid is a multi-targeted kinase inhibitor primarily targeting Aurora A/B and JAK2/3. It demonstrates potent biological activity against various cancers, exhibiting IC50 values between 1 to 30 nM for its targets. AT9283 lactic acid effectively inhibits the growth and survival of multiple solid tumors in both in vitro and in vivo models, making it a valuable reagent for cancer research applications. -
BRD4 Inhibitor
BRD4 Inhibitor-18 is a potent inhibitor of the Bromodomain-containing protein 4 (BRD4), exhibiting an IC50 value of 110 nM. This compound features a hydrophobic acetylcyclopentanyl side chain and significantly reduces the proliferation of MV-4-11 leukemia cells, which are characterized by high BRD4 expression. In addition, BRD4 Inhibitor-18 promotes apoptosis and induces G0/G1 cell cycle arrest, making it a valuable tool for research into cancer therapeutics and cell cycle regulation. -
CDK6/BRD4 Inhibitor
BC13 is a selective inhibitor of CDK6 and BRD4, demonstrating IC50 values of 234 nM and 36 nM, respectively. This compound exhibits notable antiproliferative effects, facilitating cell apoptosis and inducing DNA damage in various cell lines. Additionally, BC13 has been shown to elevate reactive oxygen species (ROS) levels, making it a valuable tool for research in cancer biology and therapeutic development targeting cell cycle regulation. -
Sirtuin Inhibitor
Sirt1/2-IN-3 is a dual inhibitor of the sirtuin family, specifically targeting SIRT1 and SIRT2 with IC50 values of 1.4 μM and 2.0 μM, respectively. This compound effectively prevents the deacetylation of p53, leading to increased acetylation of both p53 and α-tubulin. Sirt1/2-IN-3 has been demonstrated to induce apoptosis and exhibit anti-proliferative effects on human leukemia cell lines, making it a valuable tool for cancer research and the study of cellular aging mechanisms. -
SIRT Inhibitor
SIRT-IN-7 is a selective inhibitor targeting the SIRT family of proteins, specifically SIRT1, SIRT2, and SIRT3. This compound enhances the acetylation and activation of the tumor suppressor protein p53, leading to the inhibition of proliferation and the induction of apoptosis and autophagy in breast cancer cells. SIRT-IN-7 demonstrates significant anti-tumor activity, making it a valuable tool for research in cancer biology and therapeutic development. -
PARP-2 Inhibitor
PARP-2-IN-3 is a potent inhibitor of PARP-2, exhibiting an IC50 of 0.07 μM. This compound effectively induces apoptosis and necrosis in cancer cells, making it a valuable tool for cancer research. Additionally, PARP-2-IN-3 demonstrates favorable pharmacokinetic properties and oral bioavailability, supporting its potential use in therapeutic applications targeting PARP-2 related pathways. -
HDAC Inhibitor
HDAC-IN-73 is a potent histone deacetylase (HDAC) inhibitor targeting HDAC1 and HDAC6, with IC50 values of 0.17 µM and 0.49 µM, respectively. Its enhanced activity against HDAC6 demonstrates a nine-fold greater potency compared to PsA, making it a valuable compound in the field of cancer research. HDAC-IN-73 exhibits significant antiproliferative effects, induces apoptosis, and triggers G2/M cell cycle arrest, positioning it as a promising candidate for investigating therapies in colon cancer and other malignancies. -
HDAC Inhibitor
HDAC-IN-96 is a selective inhibitor of histone deacetylases 1 and 2 (HDAC1/2), exhibiting IC50 values of 457.1 nM and 433.7 nM, respectively. This compound demonstrates significant cytotoxicity against various hematological tumor cell lines, including RS4;11, K562, RPMI-8226, and U266, with IC50 values between 2.11 and 5.35 μM. HDAC-IN-96 has been shown to induce apoptosis and cause S phase arrest in cancer cells, making it a valuable tool for research in hematological malignancies such as acute lymphoblastic leukemia. -
Aurora A/Aurora B/HDAC1/HDAC2 Inhibitor
Aurora kinase/HDAC-IN-1 is a potent dual inhibitor targeting Aurora A, Aurora B, HDAC1, and HDAC2. This compound promotes histone H3 acetylation, inhibits Aurora A phosphorylation and downstream signaling, and induces apoptosis through G2/M cell-cycle arrest. It demonstrates significant antiproliferative activity in colorectal cancer cells, with an IC50 of 30.2 nM in HCT-116 cells, and effectively suppresses tumor growth in HCT-116 colorectal cancer xenograft mouse models. This reagent is valuable for research in cancer biology and therapeutic application development. -
HDAC1/CDK7 Inhibitor
HDAC1/CDK7-IN-1 is a dual inhibitor targeting HDAC1 and CDK7, exhibiting IC50 values of 893 nM and 248 nM, respectively. This compound effectively inhibits the proliferation of cancer cell lines, including MDA-MB-231, MCF-7, A549, and HCT-116. Additionally, HDAC1/CDK7-IN-1 induces cell cycle arrest and apoptosis specifically in HCT-116 cells, while also disrupting their migratory capacity. These properties make it a valuable tool for cancer research, particularly in exploring therapeutic strategies that target epigenetic regulation and cell cycle dynamics. -
HDAC Inhibitor
WMJ-J-09 is a potent HDAC inhibitor with sub-nanomolar activity, exhibiting IC50 values of 7.5 nM against HDAC1 and 3.9 nM against HDAC6, along with notable activity towards HDAC2, HDAC3, and HDAC8. This compound effectively disrupts the cell cycle and promotes apoptosis in cancer cells through the LKB1-AMPK-p38MAPK-p63-survivin signaling pathway. By inhibiting HDAC enzyme activity, WMJ-J-09 leads to the acetylation of critical proteins, thus contributing to the regulation of cell death in cancer models, such as HCT116 and FaDu cells. -
HDAC Inhibitor
TH-6 is a potent inhibitor of histone deacetylases (HDACs), demonstrating IC50 values of 0.115 µM for HDAC1, 0.135 µM for HDAC2, 0.242 µM for HDAC3, 0.138 µM for HDAC6, and 2.120 µM for HDAC8. This compound effectively inhibits cell migration and invasion while promoting apoptosis and inducing cell cycle arrest in the G2/M phase. TH-6 exhibits significant anti-tumor activity, making it a valuable tool for cancer research and therapeutic studies. -
JAK2/3 Inhibitor
JAK-2/3-IN-3 is a potent inhibitor of JAK2 and JAK3, demonstrating IC50 values of 13.00 nM and 14.86 nM, respectively. It effectively inhibits the autophosphorylation of JAK2 and promotes apoptosis in a dose- and time-dependent manner. This compound is valuable for research into lymphoid malignancies and leukemia, providing insights into the role of JAK signaling pathways in these diseases. -
HDAC Inhibitor
HDAC-IN-36 is a potent HDAC (histone deacetylase) inhibitor that targets HDAC6 with an IC50 of 11.68 nM. This compound demonstrates significant biological activity by promoting apoptosis, enhancing autophagy, and inhibiting cellular migration. HDAC-IN-36 is applicable in cancer research, particularly in studies focusing on anti-tumor and anti-metastatic mechanisms in breast cancer. -
HDAC Inhibitor
Trichostatin C is an HDAC inhibitor that plays a crucial role in modulating gene expression by preventing the deacetylation of histones. This compound exhibits significant anticancer activity, inducing apoptosis and causing cell cycle arrest in the G2/M phase, making it particularly effective against lung cancer and urothelial bladder cancer. Additionally, Trichostatin C promotes differentiation in Friend leukemic cells and demonstrates antifungal properties, highlighting its potential in various research applications related to cancer biology and fungal infections. -
JMJD3/HDAC1/HDAC6 Inhibitor
JMJD3/HDAC-IN-1 is a dual inhibitor targeting both Jumonji domain-containing protein demethylase 3 (JMJD3) and histone deacetylases HDAC1 and HDAC6. With an IC50 value of 16 nM for HDAC1, this compound induces hypermethylation of histone H3K27 and hyperacetylation of H3K9, promoting apoptosis through cleavage of caspase-7 and PARP. JMJD3/HDAC-IN-1 demonstrates significant anti-cancer activity by inhibiting cell cloning, migration, and invasion, making it valuable in cancer research and therapeutic studies. -
KDM1A Inhibitor
Iadademstat is a selective inhibitor of KDM1A (LSD1) that demonstrates potent antileukemic activity. This orally active compound is particularly relevant for research involving relapsed or refractory acute myeloid leukemia. Its mechanism of action and specificity make it a valuable tool for exploring the therapeutic potential of KDM1A inhibition in hematological malignancies. -
HDAC3/p-STAT3 Inhibitor
1-Stearoyl-sn-glycero-3-phosphocholine is an inhibitor of histone deacetylase 3 (HDAC3) and the phosphorylation of signal transducer and activator of transcription 3 (p-STAT3). This compound has demonstrated the ability to induce apoptosis and exhibits significant anticancer activity in chronic myelogenous leukemia (CML) K562 cells. It serves as a valuable tool for researchers investigating the therapeutic potential of HDAC inhibitors in cancer treatment. -
SIRT7 Inhibitor
SIRT7 Inhibitor 97491 is a selective inhibitor of the SIRT7 enzyme, exhibiting an IC50 of 325 nM and effectively reducing its deacetylase activity in a dose-dependent manner. This compound enhances tumor suppression by stabilizing the p53 protein through acetylation at lysine residues K373 and K382. Additionally, SIRT7 Inhibitor 97491 promotes apoptosis via the caspase signaling pathway, making it a valuable tool for cancer research and studies focused on elucidating the role of SIRT7 in tumor progression. -
JMJD6 Inhibitor
SKLB325 is a selective inhibitor of Jumonji domain-containing 6 (JMJD6) with a binding affinity (KD) of 0.755 μM and an IC50 value of 0.7797 μM. This compound demonstrates significant antitumor activity against ovarian cancer in both in vivo and in vitro models, effectively inducing apoptosis. Additionally, SKLB325 has shown impressive efficacy in renal cell carcinoma (RCC), making it a valuable tool for cancer research and therapeutic exploration. -
JAK2 Inhibitor
Fedratinib hydrochloride hydrate is a selective, ATP-competitive inhibitor targeting the JAK2 kinase. With an IC50 of 3 nM for both JAK2 and the mutant JAK2V617F, it demonstrates significant potency. This compound exhibits 35-fold selectivity over JAK1 and 334-fold selectivity over JAK3. Fedratinib hydrochloride hydrate effectively induces apoptosis in cancer cells, making it a valuable tool for research in myeloproliferative disorders. -
Tip60 HAT Inhibitor
TH1834 is a selective inhibitor of the Tip60 (KAT5) histone acetyltransferase (HAT). This compound has demonstrated the ability to induce apoptosis and enhance DNA damage in breast cancer cells. Notably, TH1834 does not interfere with the activity of the related MOF HAT, making it a valuable tool for investigating the role of Tip60 in cancer biology and exploring potential therapeutic applications. -
KDM3B Inhibitor
PFI-90 is a selective inhibitor of the histone demethylase KDM3B, effectively inhibiting the action of PAX3-FOXO1. This compound induces apoptosis and promotes myogenic differentiation, leading to increased cell death. PFI-90 exhibits potential antitumor activity, making it a valuable tool for cellular and cancer research studies. -
STAT3/JAK Inhibitor
Brevilin A is a potent inhibitor of the STAT3/JAK signaling pathway, with an IC50 value of approximately 10.6 μM for STAT3. It exhibits anti-tumor properties and effectively inhibits the proliferation of cancer cells. Additionally, Brevilin A has been shown to induce both apoptosis and autophagy, making it a valuable tool for cancer research and therapeutic investigations. -
METTL3 Inhibitor
UZH1a is a selective inhibitor of METTL3, demonstrating an IC50 of 280 nM. This compound has shown potential for epitranscriptomic modulation, influencing various cellular processes. Additionally, UZH1a exhibits antitumor activity and serves as a valuable chemical probe for investigating the biological functions of METTL3 in research applications. -
LSD1 Inhibitor
Bomedemstat ditosylate is a potent and irreversible inhibitor of lysine-specific demethylase 1 (LSD1). By inhibiting LSD1, Bomedemstat ditosylate increases methylation levels of H3K4 and H3K9, leading to significant alterations in gene expression. This compound demonstrates notable anti-cancer properties, effectively inhibiting cancer cell proliferation and inducing apoptosis, making it a valuable tool for cancer research and therapeutic development. -
PKD/PIM2 Inhibitor
CRT0066101 is a potent and orally active inhibitor of Protein Kinase D (PKD) with IC50 values of 1 nM, 2.5 nM, and 2 nM for PKD1, PKD2, and PKD3, respectively. Additionally, it serves as an effective PIM2 inhibitor with an IC50 of approximately 135.7 nM. This compound exhibits notable anti-inflammatory activity demonstrated in LPS-induced lung injury models in mice, as well as anticancer effects, making it a valuable tool for research in cancer and inflammation-related studies. -
PAD Inhibitor
YW3-56 hydrochloride is a potent inhibitor of peptidylarginine deiminases (PADs). It is known to activate p53 target genes and modulate the ATF pathway, effectively blocking autophagic flux. Additionally, YW3-56 induces endoplasmic reticulum (ER) stress through the PERK-eIF2α-ATF4 signaling cascade while inhibiting the mTOR pathway. This compound has demonstrated potential in the treatment of triple-negative breast cancer, making it a valuable reagent for cancer research and therapeutic studies. -
PARP1 Inhibitor
Palacaparib is a potent inhibitor of PARP1, demonstrating over 8000-fold selectivity for PARP1 relative to PARP2, PARP3, PARP5a, and PARP6. It functions by selectively inhibiting PARP1 and trapping it at sites of single-strand breaks (SSBs), impeding DNA repair. This compound is investigated primarily for its anti-cancer properties, particularly in research related to HRD+ breast cancer and various advanced solid tumors. -
SIRT1/3 Inhibitor
SPC-180002 is a dual inhibitor of SIRT1 and SIRT3, exhibiting IC50 values of 1.13 μM and 5.41 μM, respectively. This compound disrupts redox homeostasis through reactive oxygen species (ROS) generation, resulting in enhanced stability of the p21 protein and consequential mitochondrial dysfunction. SPC-180002 effectively inhibits cell cycle progression and reduces cancer cell proliferation, while also activating the Nrf2 signaling pathway, making it a valuable tool for cancer research and studies on metabolic dysregulation. -
PRMT6 Inhibitor
EPZ020411 dihydrochloride is a selective inhibitor of protein arginine methyltransferase 6 (PRMT6) with a robust IC50 of 0.010 μM. This compound effectively blocks PRMT6-mediated methylation of histone H3 at arginine 2, which enhances cellular oxidative stress responses by reducing reactive oxygen species (ROS) production and inhibiting apoptosis. EPZ020411 dihydrochloride has significant implications for research in fields including neuropathic pain, colorectal cancer, ototoxicity, hearing loss, and glioblastoma. -
SIRT2/Hsp70 Inhibitor
YM-08 is a selective inhibitor of SIRT2 and Hsp70, exhibiting an IC50 of 19.9 μM for SIRT2. This compound effectively penetrates the blood-brain barrier, making it a valuable tool for studying neurodegenerative diseases and cellular stress responses. Its dual inhibitory activity allows for investigation into SIRT2 and Hsp70's roles in various biological processes and potential therapeutic applications. -
HSP70/SIRT2 Inhibitor
HSP70/SIRT2-IN-2 is a dual inhibitor targeting SIRT2 and HSP70, demonstrating an IC50 of 45.1±5.0 μM for SIRT2. This compound exhibits significant antitumor activity, making it a valuable tool for cancer research. Its ability to simultaneously inhibit these two proteins positions HSP70/SIRT2-IN-2 as a useful candidate for studies focused on tumor progression and potential therapeutic strategies. -
HDAC1/HDAC2 Inhibitor
MRLB-223 is a selective inhibitor of HDAC1 and HDAC2, demonstrating potent activity against tumor cells. It induces histone hyperacetylation and activates the intrinsic apoptotic pathway, leading to tumor cell apoptosis and degradation of Bcr-Abl in a caspase-dependent manner. Notably, MRLB-223 mediates p53-independent cell death in Bcr-Abl-expressing myeloid cells and shows efficacy in animal models of Eμ-myc lymphoma. This compound is valuable for research focusing on the mechanisms of lymphomagenesis and therapeutic strategies for Eμ-myc lymphoma. -
Polθ/PARP Inhibitor
Polθ/PARP-IN-1 is a potent dual inhibitor targeting DNA polymerase theta (Polθ) and poly (ADP-ribose) polymerase (PARP), exhibiting IC50 values of 45.6 nM and 5.4 nM, respectively. This compound demonstrates significant antiproliferative activity by inducing apoptosis and cell cycle arrest at the G2/M phase, leading to DNA damage. Polθ/PARP-IN-1 is applicable in cancer research and may contribute to therapeutic strategies targeting tumorigenesis. -
CK2/PIM1 Inhibitor
CK2/PIM1-IN-1 is a selective inhibitor targeting casein kinase 2 (CK2) and PIM1, demonstrating IC50 values of 3.787 μM and 4.327 μM, respectively. This compound is designed for research into proliferative disorders, particularly cancer, and has potential applications in studying kinase-related conditions such as inflammation, pain, vascular disorders, pathogenic infections, and certain immunological disorders. -
BRD2/BRD4 Inhibitor
KB-0118 is a selective inhibitor of the bromodomains BRD2 and BRD4, exhibiting Kd values of 36.7 μM and 47.4 μM, respectively. This orally active compound effectively inhibits the production of pro-inflammatory cytokines, such as TNF, IL-1β, and IL-23a, and selectively reduces Th17 cell differentiation. The compound modulates Th17-driven inflammatory processes through the epigenetic suppression of BRD4, leading to decreased expression of STAT3 and other target genes. KB-0118 demonstrates potential therapeutic benefits in models of inflammatory bowel disease (IBD). -
TET1 Inhibitor
NSC-370284 is a selective inhibitor of ten-eleven translocation 1 (TET1), targeting the enzymatic conversion of cytosine to 5-hydroxymethylcytosine (5hmC). This compound effectively reduces TET1 expression levels by interacting with STAT3/5 signaling pathways. NSC-370284 is valuable for research in epigenetics and may be utilized to explore the role of TET1 in various biological processes and disease states. -
AK/STAT Signaling Inhibitor
AUH-6-96 is a potent JAK/STAT signaling inhibitor that effectively reduces Unpaired-induced transcriptional activity in Drosophila cells and inhibits tyrosine phosphorylation of STAT92E. It also suppresses both constitutive and IL-6-induced phosphorylation of STAT3 while decreasing levels of tyrosine-phosphorylated JAK3. Furthermore, AUH-6-96 induces apoptosis in cancer cells by downregulating anti-apoptotic genes downstream of STAT3, selectively reducing the viability of cancer cells with dysregulated JAK/STAT signaling. This compound is relevant for research pertaining to Hodgkin's lymphoma, breast cancer, and prostate cancer. -
LSD1 Inhibitor
LSD1-IN-14 is a potent and selective inhibitor of Lysine-specific demethylase 1 (LSD1), exhibiting an IC50 of 0.89 μM. This compound effectively inhibits the proliferation of A549 lung cancer cells and THP-1 monocytic cells, while also inducing apoptosis in tumor cell lines. LSD1-IN-14 is valuable for research applications focused on understanding the role of LSD1 in cancer biology and developing potential therapeutic strategies targeting epigenetic regulation. -
HDAC Inhibitor
HDAC-IN-37 is a potent inhibitor of histone deacetylases (HDACs), demonstrating IC50 values of 0.0551 μM for HDAC1, 1.24 μM for HDAC3, 0.948 μM for HDAC8, and 34.2 μM for HDAC6. This compound effectively increases histone acetylation through a slow-off binding mechanism. Additionally, HDAC-IN-37 disrupts the transition from the G1 phase to the S phase of the cell cycle and promotes early apoptosis in various cell types, making it a valuable tool for research in cancer biology and therapeutic development. -
c-Met/HDAC Inhibitor
c-Met/HDAC-IN-3 is a dual inhibitor targeting c-Met and histone deacetylase 1 (HDAC1), exhibiting IC50 values of 12.50 nM and 26.97 nM, respectively. This compound demonstrates significant biological activity by inducing apoptosis and causing cell cycle arrest at the G2/M phase. c-Met/HDAC-IN-3 serves as a valuable tool for research in cancer biology and therapeutic development, particularly in studies focused on synergistic inhibition of oncogenic pathways. -
PARP-1/2/TNKS1/2 Inhibitor
PARP1/2/TNKS1/2-IN-1 is an inhibitor targeting PARP-1, PARP-2, TNKS1, and TNKS2, with IC50 values of 0.25 nM, 1.2 nM, 13.5 nM, and 4.15 nM, respectively. This compound demonstrates significant antitumor activity and promotes apoptosis, making it a valuable tool for research focused on cancer biology and therapeutic strategies. Its dual inhibitory action can facilitate the exploration of cellular repair mechanisms and enhance the understanding of poly(ADP-ribose) polymerases in cancer treatment. -
HDAC6 Inhibitor
HDAC6-IN-4 is a potent and selective inhibitor of histone deacetylase 6 (HDAC6), exhibiting an IC50 value of 23 nM. This compound promotes apoptosis in cancer cells and demonstrates significant antitumor efficacy while exhibiting minimal toxicity. HDAC6-IN-4 is valuable for research in cancer biology, particularly in studies focused on epigenetic regulation and therapeutic development. -
HDAC Inhibitor
HDAC-IN-31 is a selective and orally active histone deacetylase (HDAC) inhibitor, exhibiting IC50 values of 84.90 nM for HDAC1, 168.0 nM for HDAC2, 442.7 nM for HDAC3, and greater than 10,000 nM for HDAC8. This compound induces apoptosis and triggers G2/M phase cell cycle arrest, demonstrating significant antitumor efficacy. HDAC-IN-31 is applicable in research focused on diffuse large B-cell lymphoma and other cancer studies.
