- Cat. No.
- A22316
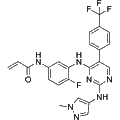
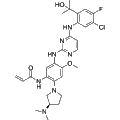
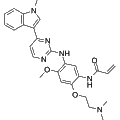
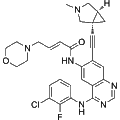
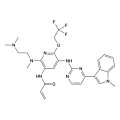
Firmonertinib (Alflutinib)
Synonyms: Furmonertinib; AST2818; Alflutinib
Alflutinib is an EGFR inhibitor that targets both EGFR activating mutations and T790M, thus leading to tumor growth inhibition.
Search by product name, CAS, target or catalog number. Press Down Arrow to reach product suggestions, Tab to move between links, and Escape to close suggestions.
EGFR and HER2 inhibitors act on two closely related receptor tyrosine kinases of the ErbB family. The epidermal growth factor receptor (EGFR, ErbB1) and human epidermal growth factor receptor 2 (HER2, ErbB2) form dimers with each other and with ErbB3 and ErbB4. Receptor activation engages the mitogen-activated protein kinase (MAPK) and phosphoinositide 3-kinase (PI3K) pathways that control cell growth. Activating EGFR mutations in non-small-cell lung cancer have made the receptor a central model in kinase inhibitor research. Gefitinib and erlotinib are reversible EGFR inhibitors, and lapatinib inhibits both EGFR and HER2. Osimertinib binds covalently to EGFR carrying the T790M resistance mutation, and neratinib irreversibly inhibits HER2 and EGFR.
Synonyms: Furmonertinib; AST2818; Alflutinib
Alflutinib is an EGFR inhibitor that targets both EGFR activating mutations and T790M, thus leading to tumor growth inhibition.
AG1557 is a specific and ATP competitive inhibitor of epidermal growth factor receptor (EGFR) tyrosine kinase, has a pIC50 value of 8.194.
SU5204, a tyrosine kinase inhibitor, has IC50s of 4 and 51.5 μM for FLK-1 (VEGFR-2) and HER2, respectively.
Synonyms: TAK-788; AP32788
Mobocertinib (TAK-788) is a potent and orally active inhibitor of EGFR and HER2 oncogenic mutants, including exon 20 insertions, with selectivity over WT EGFR. Antitumor activity.
ZD-4190 is a potent, orally available inhibitor of the vascular endothelial cell growth factor receptor 2 (VEGFR2) and of epidermal growth factor receptor (EGFR) signalling, used for the treatment of cancer.
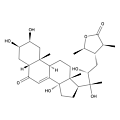
Cyasterone, a natural EGFR inhibitor, mainly isolated from Ajuga decumbens Thunb (Labiatae).
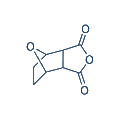
Norcantharidin (Endothall anhydride) is a synthetic anticancer compound which is a dual inhibitor for c-Met and EGFR in human colon cancers.
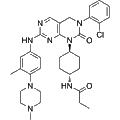
Synonyms: BLU 451
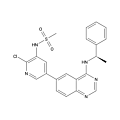
Pebezertinib (BLU-451) is an orally bioavailable epidermal growth factor receptor (EGFR) inhibitor with demonstrated central nervous system (CNS) penetration. It is specifically designed to target EGFR mutations, including exon 20 insertions, and is being investigated for the treatment of non-small cell lung cancer (NSCLC) harboring these alterations. Pebezertinib represents a promising therapeutic candidate for overcoming resistance in EGFR-driven NSCLC, particularly in cases with CNS involvement.
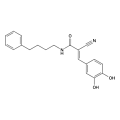
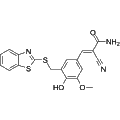
Synonyms: (E)-Tyrphostin AG 556
(E)-AG 556 is a highly selective inhibitor of epidermal growth factor receptor (EGFR) that also exhibits anti-inflammatory activity by blocking lipopolysaccharide (LPS)-induced tumor necrosis factor-alpha (TNF-α) production. Its dual functionality makes it a valuable compound for investigating EGFR-driven signaling pathways as well as inflammation-related mechanisms.
Synonyms: DZD9008
Sunvozertinib (DZD9008) is a potent, orally active inhibitor of ErbB family kinases, including mutant forms of EGFR and HER2, as well as Bruton's tyrosine kinase (BTK). It demonstrates strong inhibitory activity against a range of clinically relevant EGFR mutations, with IC₅₀ values of 20.4 nM for EGFR exon 20 NPH insertion, 20.4 nM for EGFR exon 20 ASV insertion, 1.1 nM for EGFR L858R/T790M, and 7.5 nM for HER2 exon 20 YVMA mutation. It exhibits reduced activity against wild-type EGFR (IC₅₀ = 80.4 nM in A431 cells), supporting its selectivity for mutant forms. Sunvozertinib is being investigated as a targeted therapy for non-small cell lung cancers harboring EGFR or HER2 exon 20 alterations.
CH7233163 is a noncovalent, ATP-competitive inhibitor that selectively targets the EGFR-Del19/T790M/C797S triple mutation, a known resistance mechanism to third-generation EGFR inhibitors such as Osimertinib. It effectively inhibits EGFR phosphorylation in Del19/T790M/C797S-mutant NIH3T3 cells and demonstrates significant antitumor activity in preclinical models. CH7233163 offers a promising therapeutic strategy for overcoming resistance in EGFR-mutant non-small cell lung cancer.
BLU-945 is a potent, highly selective, reversible, and orally bioavailable tyrosine kinase inhibitor (TKI) targeting mutant forms of the epidermal growth factor receptor (EGFR). It effectively inhibits EGFR variants harboring activating mutations such as L858R or exon 19 deletions, as well as resistance-associated mutations including T790M and C797S. BLU-945 is being developed as a next-generation therapeutic agent for the treatment of EGFR-mutant non-small cell lung cancer (NSCLC), particularly in cases resistant to earlier-generation EGFR inhibitors.
Synonyms: ASK120067
Limertinib (ASK120067) is a potent and orally active third-generation epidermal growth factor receptor (EGFR) tyrosine kinase inhibitor (TKI) that selectively targets the EGFR^T790M resistance mutation with an IC₅₀ of 0.3 nM, while exhibiting reduced activity against wild-type EGFR (EGFR^WT, IC₅₀ = 6.0 nM). It is being investigated as a targeted therapy for non-small cell lung cancer (NSCLC) harboring EGFR-activating and resistance mutations.
Synonyms: Tyrphostin AG-825
AG-825 is a selective, ATP-competitive inhibitor of ErbB2 (HER2) tyrosine kinase, with an IC₅₀ of 0.35 μM. It exhibits both anticancer and anti-inflammatory activities and has been shown to significantly accelerate apoptosis in human neutrophils. AG-825 also increases β₁-adrenergic receptor (β₁AR) density, suggesting potential cardiomodulatory effects. Due to its multifaceted biological activity, AG-825 is a valuable compound for research in oncology, inflammation, and cardiovascular disease.
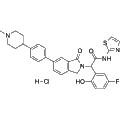
JBJ-09-063 hydrochloride is a highly potent, mutant-selective allosteric inhibitor of epidermal growth factor receptor (EGFR), specifically designed to target both TKI-sensitive and TKI-resistant EGFR mutations. It exhibits exceptionally low IC₅₀ values of: * 0.147 nM for EGFR L858R * 0.063 nM for EGFR L858R/T790M * 0.083 nM for EGFR L858R/T790M/C797S * 0.396 nM for EGFR^LT/L747S JBJ-09-063 hydrochloride effectively suppresses phosphorylation of EGFR and downstream signaling components, including Akt and ERK1/2, thereby inhibiting oncogenic signaling pathways. Its robust efficacy across a range of EGFR mutation profiles—including triple mutants that are resistant to third-generation TKIs—makes it a promising candidate for research and development in the treatment of EGFR-mutant non-small cell lung cancer (NSCLC).
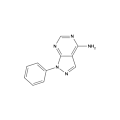
PP 3 (Compound 3) is an EGFR tyrosine kinase inhibitor with an IC50 of 2.7 μM. It targets the EGFR signaling pathway, which is critical for cell proliferation and survival in various cancers. PP 3 is a useful tool for research into EGFR-driven malignancies, such as non-small cell lung cancer and head and neck squamous cell carcinoma, enabling studies on tumor growth inhibition and potential therapeutic strategies.
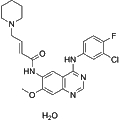
Synonyms: PF-00299804 hydrate; PF-299804 hydrate
Dacomitinib (PF-00299804) hydrate is an orally active, irreversible pan-ErbB inhibitor targeting EGFR, HER2, and HER4. It potently inhibits ErbB family signaling pathways, suppressing tumor proliferation and survival. Dacomitinib hydrate is a valuable agent for research into cancers, particularly metastatic non-small cell lung cancer (NSCLC), where it demonstrates efficacy in targeting EGFR-driven oncogenesis.
Synonyms: BPI-7711
Rezivertinib (BPI-7711) is an orally active, highly selective, and irreversible third-generation epidermal growth factor receptor (EGFR) tyrosine kinase inhibitor (TKI). It is designed to potently target both common activating EGFR mutations and the resistance-associated T790M mutation, which is a frequent cause of acquired resistance to earlier-generation TKIs. Rezivertinib also demonstrates excellent central nervous system (CNS) penetration, making it effective against brain metastases in EGFR-mutant non-small cell lung cancer (NSCLC). With its strong antitumor activity and favorable pharmacokinetic profile, Rezivertinib is a promising candidate for the treatment of EGFR-mutant NSCLC, particularly in patients with CNS involvement or T790M-driven resistance.
MTX-531 is an orally active small molecule that inhibits EGFR (IC50 = 14.7 nM) and multiple PI3K isoforms, with IC50 values of 6.4 nM (PI3Kα), 233 nM (PI3Kβ), 8.3 nM (PI3Kγ), and 1.1 nM (PI3Kδ), demonstrating potent antitumor activity. Additionally, MTX-531 functions as a weak PPARγ agonist (IC50 = 2.5 µM), which may mitigate PI3K inhibitor-induced hyperglycemia.
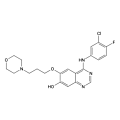
O-Desmethyl gefitinib is an active plasma metabolite of gefitinib, formed via CYP2D6-mediated metabolism. It retains EGFR inhibitory activity with an IC50 of 36 nM in subcellular assays.
Synonyms: D-0316
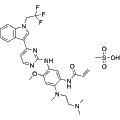
Befotertinib (D-0316) mesylate is an orally active EGFR tyrosine kinase inhibitor that suppresses tumor cell proliferation. It is primarily investigated for its potential in treating EGFR T790M-positive non-small cell lung cancer (NSCLC).
Synonyms: BDTX-1535; EGFR-IN-76
Silevertinib (BDTX-1535, EGFR-IN-76) is an orally bioavailable, blood-brain barrier-permeable, and selective EGFR inhibitor with demonstrated antitumor activity. It has shown efficacy in preclinical models of non-small cell lung cancer (NSCLC), glioblastoma patient-derived tumors, and intracranial tumor models.
JND3229 is a reversible EGFR C797S inhibitor with IC50 values of 5.8 nM for EGFR^L858R/T790M/C797S, 6.8 nM for EGFR^WT, and 30.5 nM for EGFR^L858R/T790M. It exhibits potent antiproliferative activity and effectively suppresses tumor growth in vivo, making it a valuable tool for cancer research, particularly in the context of non-small cell lung carcinoma.
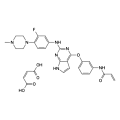
Synonyms: Abivertinib maleate; AC0010 maleate
Avitinib (Abivertinib) maleate is a third-generation, irreversible, and orally active selective EGFR inhibitor with IC50 values of 0.18 nM for both EGFR^L858R and EGFR^T790M, and 7.68 nM for wild-type EGFR. In addition to its EGFR-targeting activity, Avitinib maleate also inhibits BTK phosphorylation and induces apoptosis in mantle cell lymphoma models, demonstrating broad-spectrum anticancer efficacy.
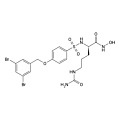
JG26 is a potent ADAM inhibitor with IC50 values of 12 nM for ADAM8, 1.9 nM for ADAM17, and 150 nM for ADAM10. It also inhibits MMP-12 with an IC50 of 9.4 nM. JG26 suppresses AngII-induced EGFR transactivation and ERK activation, upregulates ACE2 expression, inhibits CD23 shedding, and reduces SARS-CoV-2 infection. Additionally, JG26 demonstrates anti-metastatic effects in colorectal cancer and holds research potential in Hodgkin lymphoma and vascular diseases.